Most of our knowledge about bacterial functional roles in microbiomes comes from bulk measurements. Yet microbial communities are complex ecosystems in which functionally distinct bacterial subpopulations with unique transcriptional states emerge across environmental niches and from interactions with other community members. Such heterogeneous transcriptional states are inherently missed by bulk measurements. To address this gap, we developed multispecies microbial split-pool ligation meta-transcriptomics (metaSPLiT), a scalable, instrument-free single-cell RNA sequencing approach for the microbiome. Using metaSPLiT, we profiled healthy human fecal microbiomes and reconstructed 21,598 single cell transcriptomes belonging to 70 unique bacterial species.
Phenotypic heterogeneity in the human gut microbiome revealed by subspecies-resolution single-cell transcriptomics
2026-08-04 2m read
A comparison of deep multiomics profiles across ethnicity, geography, and age
2026-05-14 1m read
Despite extensive research, molecular differences in human populations and the influence of ancestry, age, geography, and diet are poorly understood. We performed comprehensive multiomics profiling (including genomics, transcriptomics, proteomics, metabolomics, lipidomics, metallomics, glycomics, and microbiomics) on samples from 322 healthy individuals of European, East Asian, and South Asian ancestry across multiple continents. We identified ethnicity-associated molecular features linked to host metabolism, autoimmune disease risk, drug metabolism, and neurodegenerative pathways. We uncovered ancestry- and geography-related molecular changes affecting metabolism, immune function, microbiome composition, and biological aging.
Cross-domain metabolic interactions link Methanobrevibacter smithii to colorectal cancer microbial ecosystems
2026-02-20 1m read
The human gut is colonized by trillions of microbes that influence the health of their human host. Whereas many bacterial species have now been linked to a variety of different diseases, the involvement of Archaea, an evolutionarily distinct group of microbes, in human disease remains elusive. By analyzing 19 independent clinical studies, we demonstrate that associations between Archaea and human diseases are widespread yet highly heterogeneous, with a pronounced and consistent enrichment of Methanobrevibacter smithii in colorectal cancer (CRC) patients. Metabolic modelling and in vitro co-culture identified distinct mutualistic interactions of M. smithii with CRC-causing bacteria such as Fusobacterium nucleatum, including metabolic enhancement. Metabolomics further reveal archaeal-derived compounds with tumor-modulating properties. Together, our results provide mechanistic insights into how the human gut archaeome may participate in CRC-associated microbial networks through metabolic cooperation with bacteria.
Integration of modern technologies to advance dietary assessment
2026-01-26 1m read
Diet is a key determinant of human and planetary health, but accurately measuring dietary intake remains challenging. Traditional self-reporting tools are imprecise, compromising our ability to accurately link diets with health outcomes. Modern technologies, including smartphone apps, image-based methods and biomarkers of food intake (BFIs), offer promise but bring their own caveats. App- and image-based methods reduce bias and reporting burden, but remain partly self-reported, and are thus prone to errors similar to those of traditional methods. Omics-based BFIs (that is, metabolites, food-related DNA or food proteins) are objective measures derived from biological samples; however, they mostly reflect recent intake, and require careful sampling alignment to estimate habitual diets. Here we discuss the drawbacks and opportunities for all dietary tools and propose strategies to integrate technologies along with multisampling for longitudinal measurements, for a new era in dietary assessment that can clarify the impact of diets, dietary components and dietary behaviour on human and planetary health.
MICOMWeb: a website for microbial community metabolic modeling of the human gut
2025-12-02 1m read
MICOMWeb is a user-friendly website for modeling microbial community metabolism in the human gut. This website tackles three constraints when generating in silico metagenome-scale metabolic models: i) the prior Python user knowledge for metabolic modeling using flux balance analysis with the MICOM Python package, ii) predefined and user-defined diets to generate ad hoc metabolic models, and iii) the high-throughput computational infrastructure required to obtain the simulated growth and metabolic exchange fluxes, using real abundance from metagenomic shotgun or 16S amplicon sequencing; we present MICOMWeb’s features to easily run in silico experiments as a functional hypothesis generator for experimental validation on three previously published databases.
Personalized Clostridioides difficile colonization risk prediction and probiotic therapy assessment in the human gut
2025-08-20 1m read
Clostridioides difficile (C. difficile) colonizes up to 40% of community-dwelling adults without causing disease but can eventually lead to infection (C. difficile infection [CDI]). There has been a lack of focus on how to prevent colonization and facilitate the successful clearance of C. difficile prior to the emergence of CDI. We show that microbial community-scale metabolic models (MCMMs) accurately predict C. difficile colonization susceptibility in vitro and in vivo, offering mechanistic insights into microbiota-specific interactions involving metabolites like succinate, trehalose, and ornithine. MCMMs reveal distinct C. difficile metabolic niches—two growth-associated and one non-growth-associated—observed across 15,204 individuals from five cohorts. We further demonstrate that MCMMs can predict personalized C. difficile growth suppression by a probiotic cocktail designed to replace fecal microbiota transplants (FMTs) for the treatment of recurrent CDI, and we identify new probiotic targets for future validation. MCMMs represent a powerful framework for predicting pathogen colonization and assessing probiotic efficacy across diverse microbiota contexts.
Metagenomic estimation of absolute bacterial biomass in the mammalian gut through host-derived read normalization
2025-07-31 2m read
Absolute bacterial biomass estimation in the human gut is crucial for understanding microbiome dynamics and host-microbe interactions. Current methods for quantifying bacterial biomass in stool, such as flow cytometry, quantitative polymerase chain reaction (qPCR), or spike-ins, can be labor-intensive, costly, and confounded by factors like water content, DNA extraction efficiency, PCR inhibitors, and other technical challenges that add bias and noise. We propose a simple, cost-effective approach that circumvents some of these technical challenges: directly estimating bacterial biomass from metagenomes using bacterial-to-host (B:H) read count ratios.
Understanding disease-associated metabolic changes in human colonic epithelial cells using the iColonEpithelium metabolic reconstruction
2025-07-03 2m read
The colonic epithelium plays a key role in the host-microbiome interactions, allowing uptake of various nutrients and driving important metabolic processes. To unravel detailed metabolic activities in the human colonic epithelium, our present study focuses on the generation of the first cell-type-specific genome-scale metabolic model (GEM) of human colonic epithelial cells, named iColonEpithelium.
GEMs are powerful tools for exploring reactions and metabolites at the systems level and predicting the flux distributions at steady state. Our cell-type-specific iColonEpithelium metabolic reconstruction captures genes specifically expressed in the human colonic epithelial cells. iColonEpithelium is also capable of performing metabolic tasks specific to the colonic epithelium. A unique transport reaction compartment has been included to allow for the simulation of metabolic interactions with the gut microbiome. We used iColonEpithelium to identify metabolic signatures associated with inflammatory bowel disease.
Metagenomic estimation of dietary intake from human stool
2025-02-18 1m read
Dietary intake is tightly coupled to gut microbiota composition, human metabolism and the incidence of virtually all major chronic diseases. Dietary and nutrient intake are usually assessed using self-reporting methods, including dietary questionnaires and food records, which suffer from reporting biases and require strong compliance from study participants. Here, we present Metagenomic Estimation of Dietary Intake (MEDI): a method for quantifying food-derived DNA in human faecal metagenomes.
We show that DNA-containing food components can be reliably detected in stool-derived metagenomic data, even when present at low abundances (more than ten reads). We show how MEDI dietary intake profiles can be converted into detailed metabolic representations of nutrient intake. MEDI identifies the onset of solid food consumption in infants, shows significant agreement with food frequency questionnaire responses in an adult population and shows agreement with food and nutrient intake in two controlled-feeding studies. Finally, we identify specific dietary features associated with metabolic syndrome in a large clinical cohort without dietary records, providing a proof-of-concept for detailed tracking of individual-specific, health-relevant dietary patterns without the need for questionnaires.
Neonates exposed to HIV but uninfected exhibit an altered gut microbiota and inflammation associated with impaired breast milk antibody function
2024-12-20 1m read
Background
Infants exposed to HIV but uninfected have altered immune profiles which include heightened systemic inflammation. The mechanism(s) underlying this phenomenon is unknown. Here, we investigated differences in neonatal gut bacterial and viral microbiome and associations with inflammatory biomarkers in plasma. Further, we tested whether HIV exposure impacts antibody-microbiota binding in neonatal gut and whether antibodies in breast milk impact the growth of commensal bacteria.
Results
Neonates exposed to HIV but uninfected (nHEU) exhibited altered gut bacteriome and virome compared to unexposed neonates (nHU). In addition, HIV exposure differentially impacted IgA-microbiota binding in neonates. The relative abundance of Blautia spp. in the whole stool or IgA-bound microbiota was positively associated with plasma concentrations of C-reactive protein. Finally, IgA from the breast milk of mothers living with HIV displayed a significantly lower ability to inhibit the growth of Blautia coccoides which was associated with inflammation in nHEU.
Elucidating human gut microbiota interactions that robustly inhibit diverse Clostridioides difficile strains across different nutrient landscapes
2024-08-28 1m read
The human gut pathogen Clostridioides difficile displays substantial inter-strain genetic variability and confronts a changeable nutrient landscape in the gut. We examined how human gut microbiota inter-species interactions influence the growth and toxin production of various C. difficile strains across different nutrient environments.
Negative interactions influencing C. difficile growth are prevalent in an environment containing a single highly accessible resource and sparse in an environment containing C. difficile-preferred carbohydrates. C. difficile toxin production displays significant community-context dependent variation and does not trend with growth-mediated inter-species interactions. C. difficile strains exhibit differences in interactions with Clostridium scindens and the ability to compete for proline. Further, C. difficile shows substantial differences in transcriptional profiles in co-culture with C. scindens or Clostridium hiranonis.
Aberrant bowel movement frequencies coincide with increased microbe-derived blood metabolites associated with reduced organ function
2024-07-16 1m read
Bowel movement frequency (BMF) directly impacts the gut microbiota and is linked to diseases like chronic kidney disease or dementia. In particular, prior work has shown that constipation is associated with an ecosystem-wide switch from fiber fermentation and short-chain fatty acid production to more detrimental protein fermentation and toxin production. Here, we analyze multi-omic data from generally healthy adults to see how BMF affects their molecular phenotypes, in a pre-disease context.
Microbial community-scale metabolic modeling predicts personalized short-chain-fatty-acid production profiles in the human gut
2024-06-24 1m read
Microbially derived short-chain fatty acids (SCFAs) in the human gut are tightly coupled to host metabolism, immune regulation and integrity of the intestinal epithelium. However, the production of SCFAs can vary widely between individuals consuming the same diet, with lower levels often associated with disease. A systems-scale mechanistic understanding of this heterogeneity is lacking. Here we use a microbial community-scale metabolic modelling (MCMM) approach to predict individual-specific SCFA production profiles to assess the impact of different dietary, prebiotic and probiotic inputs. We evaluate the quantitative accuracy of our MCMMs using in vitro and ex vivo data, plus published human cohort data.
Island biogeography theory provides a plausible explanation for why larger vertebrates and taller humans have more diverse gut microbiomes
2024-06-21 2m read
Prior work has shown a positive scaling relationship between vertebrate body size, human height, and gut microbiome alpha diversity. This observation mirrors commonly observed species area relationships (SAR) in many other ecosystems. Here, we expand these observations to several large data sets, showing that this size-diversity scaling relationship is independent of relevant covariates, like diet, body mass index, age, sex, bowel movement frequency, antibiotic usage, and cardiometabolic health markers. Island biogeography theory (IBT), which predicts that larger islands tend to harbor greater species diversity through neutral demographic processes, provides a simple mechanism for positive SARs.
Meta-analysis of the human upper respiratory tract microbiome reveals robust taxonomic associations with health and disease
2024-04-23 1m read
Background
The human upper respiratory tract (URT) microbiome, like the gut microbiome, varies across individuals and between health and disease states. However, study-to-study heterogeneity in reported case–control results has made the identification of consistent and generalizable URT-disease associations difficult.
Results
In order to address this issue, we assembled 26 independent 16S rRNA gene amplicon sequencing data sets from case–control URT studies, with approximately 2–3 studies per respiratory condition and ten distinct conditions covering common chronic and acute respiratory diseases. We leveraged the healthy control data across studies to investigate URT associations with age, sex, and geographic location, in order to isolate these associations from health and disease states.
Disease-specific loss of microbial cross-feeding interactions in the human gut
2023-10-20 1m read
Many gut microorganisms critical to human health rely on nutrients produced by each other for survival; however, these cross-feeding interactions are still challenging to quantify and remain poorly characterized. Here, we introduce a Metabolite Exchange Score (MES) to quantify those interactions. Using metabolic models of prokaryotic metagenome-assembled genomes from over 1600 individuals, MES allows us to identify and rank metabolic interactions that are significantly affected by a loss of cross-feeding partners in 10 out of 11 diseases. When applied to a Crohn’s disease case-control study, our approach identifies a lack of species with the ability to consume hydrogen sulfide as the main distinguishing microbiome feature of disease. We propose that our conceptual framework will help prioritize in-depth analyses, experiments and clinical targets, and that targeting the restoration of microbial cross-feeding interactions is a promising mechanism-informed strategy to reconstruct a healthy gut ecosystem.
Bifidobacterium infantis supplementation versus placebo in early life to improve immunity in infants exposed to HIV: a protocol for a randomized trial
2023-10-08 1m read
Infants who are born from mothers with HIV (infants who are HIV exposed but uninfected; iHEU) are at higher risk of morbidity and display multiple immune alterations compared to infants who are HIV-unexposed (iHU). Easily implementable strategies to improve immunity of iHEU, and possibly subsequent clinical health outcomes, are needed. iHEU have altered gut microbiome composition and bifidobacterial depletion, and relative abundance of Bifidobacterium infantis has been associated with immune ontogeny, including humoral and cellular vaccine responses. Therefore, we will assess microbiological and immunological phenotypes and clinical outcomes in a randomized, double-blinded trial of B. infantis Rosell®-33 versus placebo given during the first month of life in South African iHEU.
Growth phase estimation for abundant bacterial populations sampled longitudinally from human stool metagenomes
2023-09-04 1m read
Longitudinal sampling of the stool has yielded important insights into the ecological dynamics of the human gut microbiome. However, human stool samples are available approximately once per day, while commensal population doubling times are likely on the order of minutes-to-hours. Despite this mismatch in timescales, much of the prior work on human gut microbiome time series modeling has assumed that day-to-day fluctuations in taxon abundances are related to population growth or death rates, which is likely not the case. Here, we propose an alternative model of the human gut as a stationary system, where population dynamics occur internally and the bacterial population sizes measured in a bolus of stool represent a steady-state endpoint of these dynamics. We formalize this idea as stochastic logistic growth. We show how this model provides a path toward estimating the growth phases of gut bacterial populations in situ. We validate our model predictions using an in vitro Escherichia coli growth experiment. Finally, we show how this method can be applied to densely-sampled human stool metagenomic time series data. We discuss how these growth phase estimates may be used to better inform metabolic modeling in flow-through ecosystems, like animal guts or industrial bioreactors.
Coarse graining the human gut microbiome
2023-07-12 1m read
The composition of the human gut microbiome is heterogeneous across people. However, if you squint, co-abundant microbial genera emerge, accounting for much of this ecological variability. In this issue of Cell Host & Microbe, Frioux et al. provide a workflow for identifying these bacterial guilds, or “enterosignatures.”
More is Different: Metabolic Modeling of Diverse Microbial Communities
2023-03-21 1m read
Microbial consortia drive essential processes, ranging from nitrogen fixation in soils to providing metabolic breakdown products to animal hosts. However, it is challenging to translate the composition of microbial consortia into their emergent functional capacities. Community-scale metabolic models hold the potential to simulate the outputs of complex microbial communities in a given environmental context, but there is currently no consensus for what the fitness function of an entire community should look like in the presence of ecological interactions and whether community-wide growth operates close to a maximum.
Multiomic investigations of Body Mass Index reveal heterogeneous trajectories in response to a lifestyle intervention
2023-03-20 1m read
Multiomic profiling is useful in characterizing heterogeneity of both health and disease states. Obesity exerts profound metabolic perturbation in individuals and is a risk factor for multiple chronic diseases. Here, we report a global atlas of cross-sectional and longitudinal changes associated with Body Mass Index (BMI) across 1,100+ blood analytes, as well as their correspondence to host genome and fecal microbiome composition, from a cohort of 1,277 individuals enrolled in a wellness program.
Genome-microbiome interplay provides insight into the determinants of the human blood metabolome
2022-11-10 1m read
Variation in the blood metabolome is intimately related to human health. However, few details are known about the interplay between genetics and the microbiome in explaining this variation on a metabolite-by-metabolite level. Here, we perform analyses of variance for each of 930 blood metabolites robustly detected across a cohort of 1,569 individuals with paired genomic and microbiome data while controlling for a number of relevant covariates.
We find that 595 (64%) of these blood metabolites are significantly associated with either host genetics or the gut microbiome, with 69% of these associations driven solely by the microbiome, 15% driven solely by genetics and 16% under hybrid genome–microbiome control. Additionally, interaction effects, where a metabolite–microbe association is specific to a particular genetic background, are quite common, albeit with modest effect sizes. This knowledge will help to guide targeted interventions designed to alter the composition of the human blood metabolome.
Constraint-Based Reconstruction and Analyses of Metabolic Models: Open-Source Python Tools and Applications to Cancer
2022-07-07 2m read
The influence of metabolism on signaling, epigenetic markers, and transcription is highly complex yet important for understanding cancer physiology. Despite the development of high-resolution multi-omics technologies, it is difficult to infer metabolic activity from these indirect measurements. Fortunately, genome-scale metabolic models and constraint-based modeling provide a systems biology framework to investigate the metabolic states and define the genotype-phenotype associations by integrations of multi-omics data. Constraint-Based Reconstruction and Analysis (COBRA) methods are used to build and simulate metabolic networks using mathematical representations of biochemical reactions, gene-protein reaction associations, and physiological and biochemical constraints. These methods have led to advancements in metabolic reconstruction, network analysis, perturbation studies as well as prediction of metabolic state. Most computational tools for performing these analyses are written for MATLAB, a proprietary software. In order to increase accessibility and handle more complex datasets and models, community efforts have started to develop similar open-source tools in Python. To date there is a comprehensive set of tools in Python to perform various flux analyses and visualizations; however, there are still missing algorithms in some key areas. This review summarizes the availability of Python software for several components of COBRA methods and their applications in cancer metabolism. These tools are evolving rapidly and should offer a readily accessible, versatile way to model the intricacies of cancer metabolism for identifying cancer-specific metabolic features that constitute potential drug targets.
Heterogeneity in statin responses explained by variation in the human gut microbiome
2022-06-10 1m read
Statins remain one of the most prescribed medications worldwide. While effective in decreasing atherosclerotic cardiovascular disease risk, statin use is associated with adverse effects for a subset of patients, including disrupted metabolic control and increased risk of type 2 diabetes.
We investigated the potential role of the gut microbiome in modifying patient responses to statin therapy across two independent cohorts (discovery n = 1,848, validation n = 991). Microbiome composition was assessed in these cohorts using stool 16S rRNA amplicon and shotgun metagenomic sequencing, respectively. Microbiome associations with markers of statin on-target and adverse effects were tested via a covariate-adjusted interaction analysis framework, utilizing blood metabolomics, clinical laboratory tests, genomics, and demographics data.
From taxonomy to metabolic output: what factors define gut microbiome health?
2022-04-23 1m read
Many studies link the composition of the human gut microbiome to aberrant health states. However, our understanding of what constitutes a ‘healthy’ gut ecosystem, and how to effectively monitor and maintain it, are only now emerging. Here, we review current approaches to defining and monitoring gut microbiome health, and outline directions for developing targeted ecological therapeutics. We emphasize the importance of identifying which ecological features of the gut microbiome are most resonant with host molecular phenotypes, and highlight certain gut microbial metabolites as potential biomarkers of gut microbiome health. We further discuss how multi-omic measurements of host phenotypes, dietary information, and gut microbiome profiles can be integrated into increasingly sophisticated host-microbiome mechanistic models that can be leveraged to design personalized interventions. Overall, we summarize current progress on defining microbiome health and highlight a number of paths forward for engineering the ecology of the gut to promote wellness.
Genomic and functional characterization of a mucosal symbiont involved in early-stage colorectal cancer
2021-10-13 1m read
Colorectal cancer is a major health concern worldwide. Growing evidence for the role of the gut microbiota in the initiation of CRC has sparked interest in approaches that target these microorganisms. However, little is known about the composition and role of the microbiota associated with precancerous polyps. Here, we found distinct microbial signatures between patients with and without polyps and between polyp subtypes using sequencing and culturing techniques. We found a correlation between Bacteroides fragilis recovered and the level of inflammatory cytokines in the mucosa adjacent to the polyp. Additional analysis revealed that B. fragilis from patients with polyps are bft-negative, activate NF-κB through Toll-like receptor 4, induce a pro-inflammatory response, and are enriched in genes associated with LPS biosynthesis. This study provides fundamental insight into the microbial microenvironment of the pre-neoplastic polyp by highlighting strain-specific genomic and proteomic differences, as well as more broad compositional differences in the microbiome.
Lettuce (Lactuca sativa) productivity influenced by microbial inocula under nitrogen-limited conditions in aquaponics
2021-02-23 2m read
The demand for food will outpace productivity of conventional agriculture due to projected growth of the human population, concomitant with shrinkage of arable land, increasing scarcity of freshwater, and a rapidly changing climate. While aquaponics has potential to sustainably supplement food production with minimal environmental impact, there is a need to better characterize the complex interplay between the various components (fish, plant, microbiome) of these systems to optimize scale up and productivity. Here, we investigated how the commonly-implemented practice of continued microbial community transfer from pre-existing systems might promote or impede productivity of aquaponics. Specifically, we monitored plant growth phenotypes, water chemistry, and microbiome composition of rhizospheres, biofilters, and fish feces over 61-days of lettuce (Lactuca sativa var. crispa) growth in nitrogen-limited aquaponic systems inoculated with bacteria that were either commercially sourced or originating from a pre-existing aquaponic system. Lettuce above- and below-ground growth were significantly reduced across replicates treated with a pre-existing aquaponic system inoculum when compared to replicates treated with a commercial inoculum. Reduced productivity was associated with enrichment in specific bacterial genera in plant roots, including Pseudomonas, following inoculum transfer from pre-existing systems. Increased productivity was associated with enrichment of nitrogen-fixing Rahnella in roots of plants treated with the commercial inoculum. Thus, we show that inoculation from a pre-existing system, rather than from a commercial inoculum, is associated with lower yields. Further work will be necessary to test the putative mechanisms involved.
Gut microbiome pattern reflects healthy ageing and predicts survival in humans
2021-02-18 1m read
The gut microbiome has important effects on human health, yet its importance in human ageing remains unclear. In the present study, we demonstrate that, starting in mid-to-late adulthood, gut microbiomes become increasingly unique to individuals with age.
We leverage three independent cohorts comprising over 9,000 individuals and find that compositional uniqueness is strongly associated with microbially produced amino acid derivatives circulating in the bloodstream. In older age (over ~80 years), healthy individuals show continued microbial drift towards a unique compositional state, whereas this drift is absent in less healthy individuals.
Progressive shifts in the gut microbiome reflect prediabetes and diabetes development in a treatment-naive Mexican cohort
2021-01-08 2m read
Type 2 diabetes (T2D) is a global epidemic that affects more than 8% of the world’s population and is a leading cause of death in Mexico. Diet and lifestyle are known to contribute to the onset of T2D. However, the role of the gut microbiome in T2D progression remains uncertain. Associations between microbiome composition and diabetes are confounded by medication use, diet, and obesity. Here we present data on a treatment-naive cohort of 405 Mexican individuals across varying stages of T2D severity. Associations between gut bacteria and more than 200 clinical variables revealed a defined set of bacterial genera that were consistent biomarkers of T2D prevalence and risk. Specifically, gradual increases in blood glucose levels, beta cell dysfunction, and the accumulation of measured T2D risk factors was correlated with the relative abundances of four bacterial genera. In a cohort of 25 individuals, T2D treatment - predominantly metformin - reliably returned the microbiome to the normoglycemic community state. Deep clinical characterization allowed us to broadly control for confounding variables, indicating that these microbiome patterns were independent of common T2D comorbidities, like obesity or cardiovascular disease. Thus, our work provides the first solid evidence for a direct link between the gut microbiome and T2D in a critically high-risk population. Whether or not these T2D-associated changes in the gut contribute to the etiology of T2D or its comorbidities remains to be seen.
Baseline Gut Metagenomic Functional Gene Signature Associated with Variable Weight Loss Responses following a Healthy Lifestyle Intervention in Humans
2021-01-05 1m read
Recent human feeding studies have shown how the baseline taxonomic composition of the gut microbiome can determine responses to dietary interventions, but the exact functional determinants underlying this phenomenon remain unclear.
In this study, we set out to better understand interactions between baseline BMI, metabolic health, diet, gut microbiome functional profiles, and subsequent weight changes in a human cohort that underwent a healthy lifestyle intervention. Overall, our results suggest that the microbiota may influence host weight loss responses through variable bacterial growth rates, dietary energy harvest efficiency, and immunomodulation.
Antimicrobial Peptide against Mycobacterium Tuberculosis That Activates Autophagy Is an Effective Treatment for Tuberculosis
2020-11-09 1m read
Mycobacterium tuberculosis (MTB) is the principal cause of human tuberculosis (TB), which is a serious health problem worldwide. The development of innovative therapeutic modalities to treat TB is mainly due to the emergence of multi drug resistant (MDR) TB. Autophagy is a cell-host defense process. Previous studies have reported that autophagy-activating agents eliminate intracellular MDR MTB. Thus, combining a direct antibiotic activity against circulating bacteria with autophagy activation to eliminate bacteria residing inside cells could treat MDR TB. We show that the synthetic peptide, IP-1 (KFLNRFWHWLQLKPGQPMY), induced autophagy in HEK293T cells and macrophages at a low dose (10 μM), while increasing the dose (50 μM) induced cell death; IP-1 induced the secretion of TNFα in macrophages and killed Mtb at a dose where macrophages are not killed by IP-1. Moreover, IP-1 showed significant therapeutic activity in a mice model of progressive pulmonary TB. In terms of the mechanism of action, IP-1 sequesters ATP in vitro and inside living cells. Thus, IP-1 is the first antimicrobial peptide that eliminates MDR MTB infection by combining four activities: reducing ATP levels, bactericidal activity, autophagy activation, and TNFα secretion.
MEMOTE for standardized genome-scale metabolic model testing
2020-03-02 1m read
Reconstructing metabolic reaction networks enables the development of testable hypotheses of an organism’s metabolism under different conditions1. State-of-the-art genome-scale metabolic models (GEMs) can include thousands of metabolites and reactions that are assigned to subcellular locations. Gene–protein–reaction (GPR) rules and annotations using database information can add meta-information to GEMs. GEMs with metadata can be built using standard reconstruction protocols2, and guidelines have been put in place for tracking provenance and enabling interoperability, but a standardized means of quality control for GEMs is lacking3. Here we report a community effort to develop a test suite named MEMOTE (for metabolic model tests) to assess GEM quality.
MICOM: metagenome-scale modeling to infer metabolic interactions in the microbiota
2020-01-21 2m read
Compositional changes in the gut microbiota have been associated with a variety of medical conditions such as obesity, Crohn’s disease, and diabetes. However, connecting microbial community composition to ecosystem function remains a challenge. Here, we introduce MICOM, a customizable metabolic model of the human gut microbiome. By using a heuristic optimization approach based on L2 regularization, we were able to obtain a unique set of realistic growth rates that corresponded well with observed replication rates.
Reproducible, interactive, scalable and extensible microbiome data science using QIIME 2
2019-07-24 1m read
Rapid advances in DNA-sequencing and bioinformatics technologies in the past two decades have substantially improved understanding of the microbial world. This growing understanding relates to the vast diversity of microorganisms; how microbiota and microbiomes affect disease and medical treatment; how microorganisms affect the health of the planet; and the nascent exploration of the medical, forensic, environmental and agricultural applications of microbiome biotechnology.
Much of this work has been driven by marker-gene surveys (for example, bacterial/archaeal 16S rRNA genes, fungal internal-transcribed-spacer regions and eukaryotic 18S rRNA genes), which profile microbiota with varying degrees of taxonomic specificity and phylogenetic information. The field is now transitioning to integrate other data types, such as metabolite, metaproteome or metatranscriptome profiles.
Use and abuse of correlation analyses in microbial ecology
2019-06-03 1m read
Correlation analyses are often included in bioinformatic pipelines as methods for inferring taxon–taxon interactions. In this perspective, we highlight the pitfalls of inferring interactions from covariance and suggest methods, study design considerations, and additional data types for improving high-throughput interaction inferences. We conclude that correlation, even when augmented by other data types, almost never provides reliable information on direct biotic interactions in real-world ecosystems. These bioinformatically inferred associations are useful for reducing the number of potential hypotheses that we might test, but will never preclude the necessity for experimental validation.
Non-responder phenotype reveals apparent microbiome-wide antibiotic tolerance in the murine gut
2019-04-06 2m read
Broad spectrum antibiotics cause both transient and lasting damage to the ecology of the gut microbiome. Antibiotic-induced loss of gut bacterial diversity has been linked to susceptibility to enteric infections. Prior work on subtherapeutic antibiotic treatment in humans and non-human animals has suggested that entire gut communities may exhibit tolerance phenotypes. In this study, we validate the existence of these community tolerance phenotypes in the murine gut and explore how antibiotic treatment duration or a diet enriched in antimicrobial phytochemicals might influence the frequency of this phenotype.
Distinct microbes, metabolites, and ecologies define the microbiome in deficient and proficient mismatch repair colorectal cancers
2019-04-06 2m read
Background
Links between colorectal cancer (CRC) and the gut microbiome have been established, but the specific microbial species and their role in carcinogenesis remain an active area of inquiry. Our understanding would be enhanced by better accounting for tumor subtype, microbial community interactions, metabolism, and ecology.
Methods
We collected paired colon tumor and normal-adjacent tissue and mucosa samples from 83 individuals who underwent partial or total colectomies for CRC. Mismatch repair (MMR) status was determined in each tumor sample and classified as either deficient MMR (dMMR) or proficient MMR (pMMR) tumor subtypes. Samples underwent 16S rRNA gene sequencing and a subset of samples from 50 individuals were submitted for targeted metabolomic analysis to quantify amino acids and short-chain fatty acids. A PERMANOVA was used to identify the biological variables that explained variance within the microbial communities. dMMR and pMMR microbial communities were then analyzed separately using a generalized linear mixed effects model that accounted for MMR status, sample location, intra-subject variability, and read depth. Genome-scale metabolic models were then used to generate microbial interaction networks for dMMR and pMMR microbial communities. We assessed global network properties as well as the metabolic influence of each microbe within the dMMR and pMMR networks.
Synthesis of multi-omic data and community metabolic models reveals insights into the role of hydrogen sulfide in colon cancer
2019-04-06 1m read
Multi-omic data and genome-scale microbial metabolic models have allowed us to examine microbial communities, community function, and interactions in ways that were not available to us historically. Now, one of our biggest challenges is determining how to integrate data and maximize data potential. Our study demonstrates one way in which to test a hypothesis by combining multi-omic data and community metabolic models. Specifically, we assess hydrogen sulfide production in colorectal cancer based on stool, mucosa, and tissue samples collected on and off the tumor site within the same individuals. 16S rRNA microbial community and abundance data were used to select and inform the metabolic models. We then used MICOM, an open source platform, to track the metabolic flux of hydrogen sulfide through a defined microbial community that either represented on-tumor or off-tumor sample communities. We also performed targeted and untargeted metabolomics, and used the former to quantitatively evaluate our model predictions. A deeper look at the models identified several unexpected but feasible reactions, microbes, and microbial interactions involved in hydrogen sulfide production for which our 16S and metabolomic data could not account. These results will guide future in vitro, in vivo, and in silico tests to establish why hydrogen sulfide production is increased in tumor tissue.
Personalized Prediction of Proliferation Rates
2017-01-19 2m read
Cancer is a complex disease and manifests in many different forms. In fact, when speaking about cancer we are probably speaking about thousand different diseases and not one.
The high level of heterogeneity between and across different cancer subtypes requires large amounts of data to study them. Luckily, we do have large data sets today. However, the kind of knowledge we can extract from those data sets depends a lot on where we got the data from. For cancer there are two major classes of data sources: lab-maintained cell lines and biopsies taken directly from a patient’s tumor. Both can provide distinct information as summarized in the following slide.
Going from metabolites to affected enzymes in cancer
2016-06-21 2m read
When studying cancer or any disease one of the things we are interested in are the alterations that cause the disease. By now, we have quite some arsenal to study genomic aberrations, however assigning those to a specific phenotype is not trivial. This is particularly true for changes affecting metabolism, since there is a myriad of regulation events that take place after gene expression and which drive metabolism.
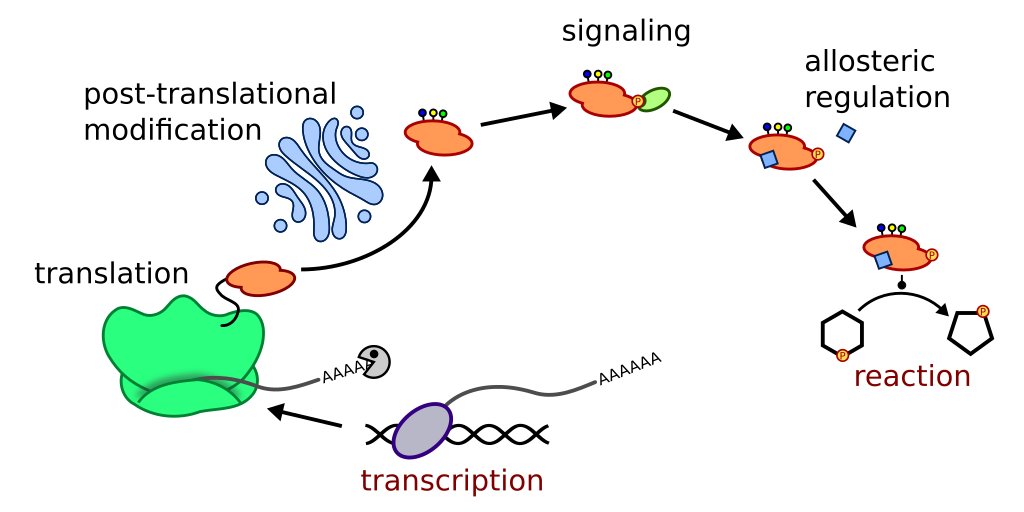
The image above shows just an example of events that can happen between translation of an enzyme gene until it will finally catalyze a reaction. Metabolome data is naturally closer to the phenotype, however it usually does not give us information on which alterations caused the observed phenotype.
Building multifunctional peptides by compatible function
2016-04-21 1m read
Artificially designed small peptides are currently quite interesting for medical research since they provide a way to target specific cellular activities. In many cases one wants to combine several functions into a single peptide that is as small as possible. This is problematic as the activity of the peptide is often lost when combining several functions.
In our article we propose a design strategy based on “compatible function”, meaning functions that require similar physio-chemical properties of the peptide. Starting from a peptide template already containing two compatible functions (DNA-binding and pheromone activity) we use a combination of Machine learning and optimization to add two additional functions (cell penetration and antibacterial) for the entire peptide by adding only a few amino acids. The resulting peptides are all very active and provide antibacterial activity on the level of Ampicillin with one peptide being even being much more efficient than Ampicillin.
How yeast creates their own signaling landscape
2015-08-28 1m read
Signaling in yeast is often used as a blueprint for human signaling pathways since its general way of function is close to what we can observe in human. This is not only true for signaling pathways within a single yeast cell but also for the ways yeast cells communicate with each other.
During the mating of yeast, individual cells communicate their position using distinct pheromones. However, at the same time they also secrete a protein that destroys those very pheromones and thos paradoxically counteracts this signaling. Using a strategy that allowed us to to visualize the pheromone landscape from fuorescence microscope images without interference, we were able to show that the interplay between creation and destruction of the pheromone allows for a much more informative signaling landscape.
CPPs and CAPs: Two sides of the same coin
2014-03-25 1m read
Cell penetrating peptides (CPP) and cationic antibacterial peptides (CAP) have similar physicochemical properties and yet it is not understood how such similar peptides display different activities. To address this question, we used Iztli peptide 1 (IP-1) because it has both CPP and CAP activities. Combining experimental and computational modeling of the internalization of IP-1, we show it is not internalized by receptor-mediated endocytosis, yet it permeates into many different cell types, including fungi and human cells. We also show that IP-1 makes pores in the presence of high electrical potential at the membrane, such as those found in bacteria and mitochondria. These results provide the basis to understand the functional redundancy of CPPs and CAPs.
Immunogenic variety and the Golden Agers
2012-11-30 2m read
The immune system protects us from foreign substances or pathogens by generating specific antibodies. The variety of immunoglobulin (Ig) paratopes for antigen recognition is a result of the V(D)J rearrangement mechanism, while a fast and efficient immune response is mediated by specific immunoglobulin isotypes obtained through class switch recombination (CSR). To get a better understanding on how antibody-based immune protection works and how it changes with age, the interdependency between these two parameters need to be addressed. Here, we have performed an in depth analysis of antibody repertoires of 14 healthy donors representing different gender and age groups. For this task, we developed a unique pyrosequencing approach, which is able to monitor the expression levels of all immunoglobulin V(D)J recombinations of all isotypes including subtypes in an unbiased and quantitative manner. Our results show that donors have individual immunoglobulin repertoires and cannot be clustered according to V(D)J recombination patterns, neither by age nor gender. However, after incorporating isotype-specific analysis and considering CSR information into hierarchical clustering the situation changes. For the first time the donors cluster according to age and separate into young adults and elderly donors (>50). As a direct consequence, this clustering defines the onset of immune senescence at the age of fifty and beyond. The observed age-dependent reduction of CSR ability proposes a feasible explanation why reduced efficacy of vaccination is seen in the elderly and implies that novel vaccine strategies for the elderly should include the “Golden Agers”.
Ensuring low noise in the yeast cell cycle
2011-10-07 2m read
The budding yeast genome comprises roughly 6000 genes generating a number of about 10 000 mRNA copies, which gives a general estimation of 1-2 mRNA copies generated per gene. What does this observation implicate for cellular processes and their regulation? Whether the number of mRNA molecules produced is important for setting the amount of proteins implicated in a particular function is at present unknown. In this context, we studied cell cycle control as one of the highly fine tuned processes that guarantee the precise timing of events essential for cell growth. Here, we developed a stochastic model that addresses the effect of varying the mRNA amount of Sic1, inhibitor of the Cdk1-Clb5 kinase activity, and the resulting noise on Sic1/Clb5 balance at the G1/S transition. We considered a range of SIC1 transcripts number according to our experimental data derived from the MS2 mRNA tagging system. Computational simulation revealed that an increased amount of SIC1 mRNAs lead to an amplified dispersion of Sic1 protein levels, suggesting mRNA control being critical to set timing of Sic1 downregulation and, therefore, S phase onset. Moreover, Sic1/Clb5 balance is strongly influenced by Clb5 production in both daughter and mother cells in order to maintain the characteristic time of S phase entry overall the population. Furthermore, CLB5 mRNA molecules calculated to reproduce temporal dynamics of Sic1 and Clb5 for daughter and mother cells agree with recent data obtained from more complex networks. Thus, the results presented here provide novel insights into the influence that the mRNA amount and, indirectly, the transcription process exploit on cell cycle progression.
What influences DNA replication rate in budding yeast?
2010-04-27 2m read
DNA replication begins at specific locations called replication origins, where helicase and polymerase act in concert to unwind and process the single DNA filaments. The sites of active DNA synthesis are called replication forks. The density of initiation events is low when replication forks travel fast, and is high when forks travel slowly. Despite the potential involvement of epigenetic factors, transcriptional regulation and nucleotide availability, the causes of differences in replication times during DNA synthesis have not been established satisfactorily, yet.